Sponsored Content by SCIEXReviewed by Olivia FrostOct 7 2026
Using the ZT Scan DIA 3.0 on the ZenoTOF 8600 system provides robust protein identification and quantitation from complex lysate digests at a throughput of up to 500 samples per day (SPD).
The precise control of sliding Q1 precursor isolation window widths (down to 1 Da) enhances quantitative accuracy and enables optimization of methods to balance proteome depth and throughput across a wide range of analytical needs.
A total of 5793 protein groups and 60,791 precursors were identified from 50 ng of a commercial human lysate digest using ZT Scan DIA 3.0 on the ZenoTOF 8600 system. This process was performed at 500 SPD, with minimal reduction in terms of total and quantifiable protein groups and precursors noted across throughputs from 60–500 SPD.
The results presented here highlight the capabilities of ZT Scan DIA 3.0 on the ZenoTOF 8600 system as a robust platform for reliable, deep protein biomarker identification and quantitation, even in large-scale cohorts.
Key high-throughput proteomics features utilizing ZT Scan DIA 3.0 on the ZenoTOF 8600 system
This approach offers a range of benefits.
Deep protein identification across varied throughput conditions
ZT Scan DIA 3.0 was found to provide deep proteome profiling from 50 ng of commercial K562 digest, successfully identifying 9268 protein groups at 60 SPD and 5793 protein groups at 500 SPD.
Advanced quantitative precision and performance
Using ZT Scan DIA 3.0, it was possible to quantify 4579 protein groups at 500 SPD and 7345 protein groups at 60 SPD from 50 ng of K562 digest, yielding median CVs of ≤7% across all throughputs for all identified protein groups.
Ultra-selective DIA supporting deeper proteome coverage
ZT Scan DIA 3.0 supports sliding Q1 isolation window widths as low as 1 Th, minimizing spectral interference from co-isolated precursors whilst also adding a high-order fourth dimension of precursor-to-fragment relationship to preserve or increase quantitative accuracy and identification confidence.
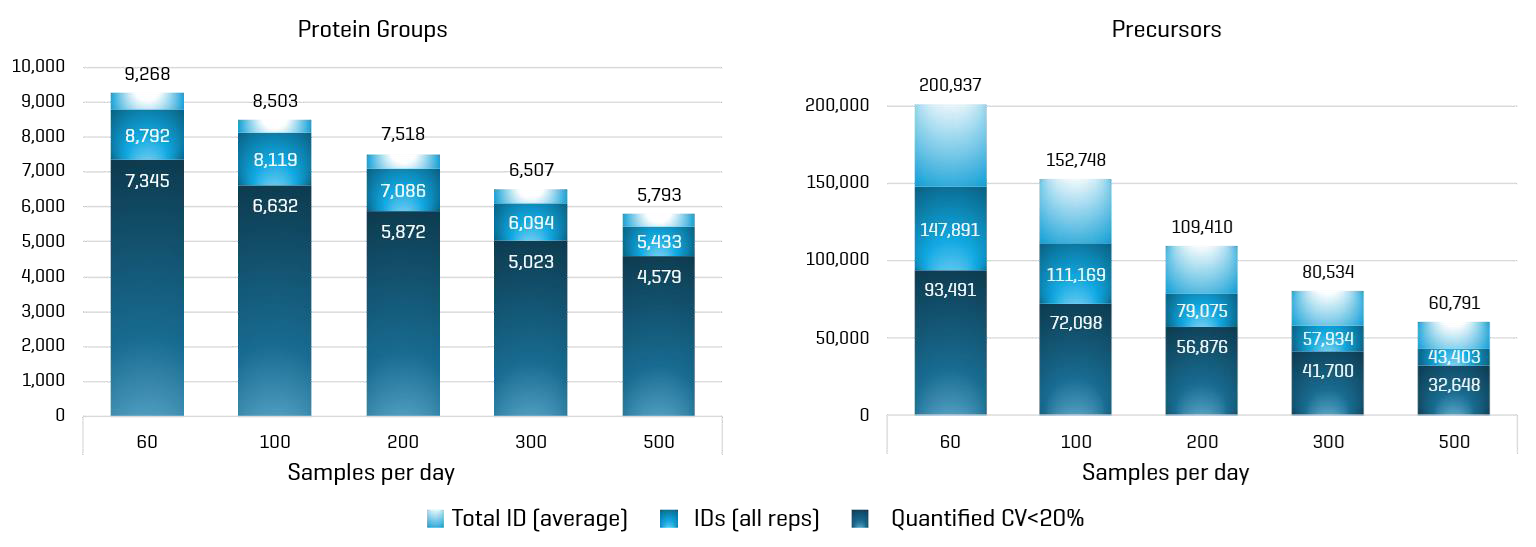
Figure 1. Protein groups and precursors identified (average IDs across 4 replicates, and IDs in all replicates) and quantified from 50 ng of K562 digest. Data-independent acquisition was performed on a ZenoTOF 8600 system using optimized ZT Scan DIA 3.0 methods at various samples-per-day (SPD) throughputs. Image Credit: SCIEX
High-throughput high-resolution mass spectrometry (HRMS) workflows are becoming increasingly essential as discovery proteomics continues to underpin faster decisions, larger studies, and broader biological insight.
Increasing throughput remains challenging, however, with shorter chromatographic separations resulting in increased precursor co-elution, compressed peptide peaks, and increased challenges in maintaining both quantitative robustness and proteome depth.
It is important to preserve proteome depth at higher throughput because deeper coverage strengthens pathway-level biological interpretation, expands access to low-abundance proteins, and increases the likelihood of detecting biologically or clinically relevant biomarkers across large sample cohorts.
These constraints can also increase interference and spectral complexity in DIA workflows, limiting the capacity for confident identification and quantitation in complex samples.
The ZenoTOF 8600 system helps address these challenges through advanced duty-cycle efficiency and ion transmission, supporting sensitive, high-confidence analysis and broad proteome coverage, even in demanding high-throughput settings.1
ZT Scan DIA was launched by SCIEX in 2024.3 This method leverages a continuously scanning quadrupole window for precursor isolation alongside a Zeno trap-activated MS/MS, facilitating higher qualitative and quantitative performance versus Zeno SWATH DIA and other traditional discrete-window DIA methods.
This acquisition method was expanded to ZT Scan DIA 2.0 in 2025, accommodating a broader mass range and enabling improved flexibility in terms of method optimization.2
The development of ZT Scan DIA 2.0 allowed users to customize Q1 window widths, but ZT Scan DIA 3.0 further extended this potential for customization, adding valuable flexibility for high-throughput studies that require maintaining depth while increasing sample turnover to maximize biological insight per unit time.
ZT Scan DIA 3.0 supports Q1 scan widths as low as 1 Th, further lowering DIA MS/MS spectra’s chimeric nature. Not only are the resulting spectra easier to interpret, but they also offer more accurate, reliable quantitation by minimizing interference stemming from neighboring precursor fragments.
This is especially important when working with complex samples and high-throughput biomarker studies, where it is important to preserve both proteome depth and quantitative precision to ensure that broad discovery can be successfully translated into confident, scalable measurement.
Users can now customize the Q1 window width, allowing researchers to effectively dial in selectivity and optimize acquisition methods to achieve the optimum balance of proteome quantitation, coverage, and sample throughput.
Methods
Sample preparation
Human K562 lysate tryptic digest was purchased from Promega before being diluted in water containing 0.1% formic acid.
Chromatography
A Vanquish Neo LC system (Thermo Scientific, USA) was used for chromatographic separations. This system utilized an Evosep EV1107 (4 cm x 0.15 mm C18 1.9 µm) or EV1109 (8 cm x 0.15 mm C18 1.5 µm) column. Mobile phase A was comprised of water and 0.1% formic acid, while mobile phase B was comprised of acetonitrile with 0.1% formic acid.
Linear gradients from 1% to 30% B were used for the different sample-per-day throughput methods, with a column wash performed at 99% B. Table 1 summarizes gradient times, flow rates, and other parameters. Four replicate injections of 50 ng K562 lysate digest were performed for each LC-MS method. The injection volume was set to 1 µL, and the column temperature to 45 °C.
Mass spectrometry
A ZenoTOF 8600 system using the microflow probe was used to perform sample analysis.
ZT Scan DIA 3.0 methods were optimized for each specific throughput, maintaining an MS/MS accumulation time of 3.3 ms and ensuring approximately five scans per chromatographic peak to achieve optimal quantitation.
Table 2 lists ion source and MS method parameters for the different throughput methods. Collision-induced dissociation fragmentation was used with the SCIEX OS software default dynamic collision energy calculation for peptides based on charge state 2. Zeno trap pulsing was enabled for all data acquisition.
Data processing
Data was processed using version 13.5 of the PEAKS Studio software using a K562/HeLa spectral library and the previously described DIA search settings.4 It should be noted that only replicate data files for a specific loading/experiment were searched together.
Calculations were performed to determine the number of quantifiable protein groups and precursors with a CV ≤20% for those identified across all four replicate injections.
Table 1. LC parameter settings used for the different SPD methods. Source: SCIEX
Throughput
(SPD) |
Column
length
(cm) |
Gradient time
(minutes) |
Flow rate
(μL/min) |
Column-wash time (minutes) |
Column-wash flow rate
(μL/min) |
MS cycle
time (seconds) |
| 60 |
8 |
21 |
1.5 |
1.3 |
1.5 |
1.6 |
| 100 |
8 |
11 |
1.5 |
2.9 |
1.5 |
0.85 |
| 200 |
4 |
5.6 |
2 |
1.2 |
2 |
0.6 |
| 300 |
4 |
3.2 |
2 |
1.2 |
2 |
0.48 |
| 500 |
4 |
2.2 |
2 |
0.5 |
4 |
0.4 |
Table 2. MS parameter settings used for the ZT Scan DIA 3.0 experiments at different samples-per-day (SPD) throughput. Source: SCIEX
| Parameter |
SPD 60 |
SPD 100 |
SPD 200 |
SPD 300 |
SPD 500 |
| Curtain gas |
|
|
40 |
|
|
| CAD gas |
|
|
7 |
|
|
| Gas 1 (psi) |
10 |
10 |
15 |
15 |
15 |
| Gas 2 (psi) |
|
|
40 |
|
|
| QJET DP (V) |
|
|
20 |
|
|
| Source temperature (°C) |
180 |
180 |
225 |
225 |
225 |
| Spray voltage (V) |
|
|
4500 |
|
|
| TOF-MS mass range (Da) |
|
|
300–1500 |
|
|
TOF MS accumulation
time (ms) |
100 |
100 |
100 |
100 |
15 |
| Q1 isolation window (Da) |
1.6 |
2.8 |
4.4 |
5 |
4.1 |
| DIA precursor range (Da) |
350–1000 |
375–960 |
370–950 |
370–950 |
400–800 |
| MS/MS mass range (Da) |
|
|
140–1750 |
|
|
MS/MS accumulation
time (ms) |
|
|
3.3 |
|
|
| Cycle time (s) |
1.6 |
0.85 |
0.6 |
0.48 |
0.4 |
Scaling proteome depth and throughput with ZT Scan DIA 3.0
Figure 1 summarizes the numbers of identified and quantified protein groups and precursors, respectively. These results were obtained from 50 ng of K562 digest using ZT Scan DIA 3.0 MS methods optimized for each specific SPD throughput.
The MS/MS accumulation time was set to 3.3 ms to optimize this, with the sliding Q1 precursor isolation window width adjusted to maintain a cycle time low enough to achieve precise quantitation via an average of five scans per chromatographic peak.
Figure 1 also shows that the narrower isolation widths utilized in the lower-throughput methods offered the highest numbers of identified and quantifiable proteins. However, throughput could be increased more than eightfold with only a minimal decrease in the number of proteins identified and quantified.
A total of 5793 protein groups were identified at 500 SPD, with 4579 of these quantifiable at ≤20% CV. A total of 60,791 precursors were also identified, with 32,648 determined to be quantifiable across all replicate injections.
Figures 2 and 3 illustrate the % CV distribution for both protein groups and precursors, respectively, across the various throughput methods. The low median %CVs highlight the ZenoTOF 8600 system with ZT Scan DIA 3.0 acquisition’s precision and reproducibility, even when operating at very high throughput.
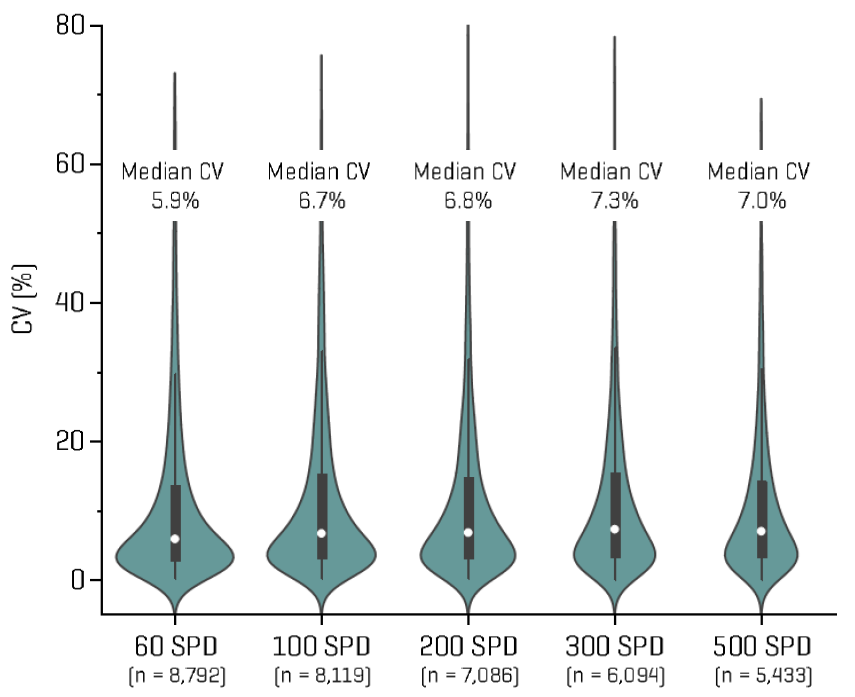
Figure 2. Violin plots of the %CV distributions for the protein groups identified (IDs in all replicates) with various throughput methods. Image Credit: SCIEX
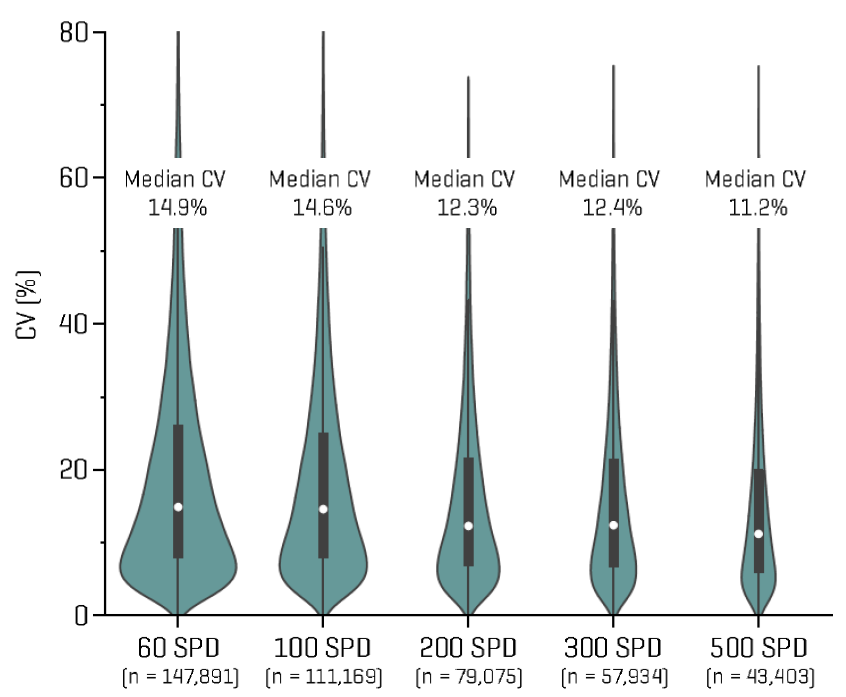
Figure 3. Violin plots of the %CV distributions for the identified precursors (IDs across all replicates) using various throughput methods. Image Credit: SCIEX
Conclusions
Rather than simply focusing on speed, the goal in high-throughput workflows is depth normalized to instrument time. The results presented here show that high-throughput discovery proteomics via ZT Scan DIA 3.0 no longer necessitates the same trade-off between data quality and speed.
This approach enables faster, large-scale studies with reproducible quantitation and robust proteome coverage.
Leveraging ZT Scan DIA 3.0 on the ZenoTDF 8600 system underpins scalable method optimization across a range of high-throughput conditions, utilizing Q1 scanning isolation widths from 1.6 to 5.0 Th to balance selectivity, proteome depth, and quantitative performance.
ZT Scan DIA 3.0 preserved deep proteome coverage for identified protein groups across the 60–500 SPD throughput range, offering a potential increase in throughput of more than eightfold (5793 versus 9268 at 60 SPD).
It was noted that increasing throughput from 60 to 500 SPD allowed broad quantifiable proteome depth to be maintained, with 4579 quantifiable protein groups at CV ≤20% across four technical replicates versus 7345 at 60 SPD.
It was also observed that narrow scanning Q1 isolation windows down to 1 Th were able to limit spectral interference and support the preservation of proteome depth while delivering reproducible precursor- and protein-level quantitation across a range of acquisition speeds.
ZT Scan DIA 3.0 helps large-cohort and translational studies by increasing sample turnover with deep biological insights. This represents a scalable path from deep discovery proteomics to high-throughput quantitative workflows.
References and further reading
- SCIEX (2020). Achieving deep quantitative proteome coverage from sub-nanogram sample loadings using Whisper Zoom and Zeno SWATH DIA on the ZenoTOF 8600 system. Available at: https://sciex.com/tech-notes/life-science-research/proteomics/achieving-deep-quantitative-proteome-coverage-from-sub-nanogram-sample-using-whisperzoom-zeno-swath-dia-on-the-zenotof-8600.
- SCIEX (2020). Advances in high-throughput quantitative proteomics powered by high-sensitivity data-independent acquisition on the ZenoTOF 8600 system. Available at: https://sciex.com/tech-notes/life-science-research/proteomics/high-throughput-quantitative-proteomics-powered-by-high-sensitivity-dia-on-the-zenotof-8600-system.
- SCIEX (2017). Continuing the data independent acquisition (r)evolution: Introducing ZT Scan DIA for quantitative proteomics. Available at: https://sciex.com/technology/zt-scan-dia/continuing-the-data-independent-acquisition-revolution-introducing-zt-scan-dia-for-quantitative-proteomics.
- SCIEX (2025). Unlocking deeper DIA performance with empirical spectral libraries generated using ZT Scan DIA 3.0 on the ZenoTOF 8600 system. Available at: https://sciex.com/tech-notes/life-science-research/proteomics/unlocking-deeper-dia-performance-with-empirical-spectral-libraries-generated-using-zt-scan-dia-3-0-on-the-zenotof-8600-system.
Acknowledgments
Produced from materials originally authored by Jason Causon, Patrick Pribil, and Remco van Soest from SCIEX.
About SCIEX
SCIEX's mission is to deliver solutions for the precision detection and quantitation of molecules, empowering their customers to protect and advance the wellness and safety of all.
SCIEX has led the field of mass spectrometry for 50 years. From the moment it launched the first ever commercially successful triple quad in 1981, it has developed groundbreaking technologies and solutions that influence life-changing research and outcomes.
Today, as part of the Danaher family of global life science and technology innovators, the company continues to pioneer robust solutions in mass spectrometry and capillary electrophoresis. The company does not just develop products. It is what it does together with its customers that sets it apart. That’s why thousands of life science experts around the world choose SCIEX to get the answers they can trust to better inform critical decisions. Decisions that positively impact lives.
The company proudly stands behind its tagline: The Power of Precision.
Sponsored Content Policy: News-Medical.net publishes articles and related content that may be derived from sources where we have existing commercial relationships, provided such content adds value to the core editorial ethos of News-Medical.net, which is to educate and inform site visitors interested in medical research, science, medical devices and treatments.